A phosphatidylserine sample that performs beautifully in a 20 kg pilot tells you almost nothing about how 500 kg of the same powder will behave on a commercial blender, a capsule filler, or a gabling line. PS is a phospholipid powder with real processing personality: it is compressible but cohesive, sensitive to heat and shear, prone to flow changes with humidity, and its assay must be built into batch calculations with the correct basis. Scale-up failures between the sample stage and commercial bulk rarely come from the ingredient being "bad" — they come from unstaged expectations: skipping a pilot at intermediate scale, treating the first commercial lot as routine production, and not scaling the documentation alongside the batch size. This guide lays out the sample-to-bulk pathway for PS finished products in OEM programs: what each stage proves, what parameters to watch, what documents the contract manufacturer must produce at each gate, and how to avoid the classic failure points that turn a promising launch into a delayed one.
Stage 0: What the Sample Stage Actually Proves — and What It Does Not
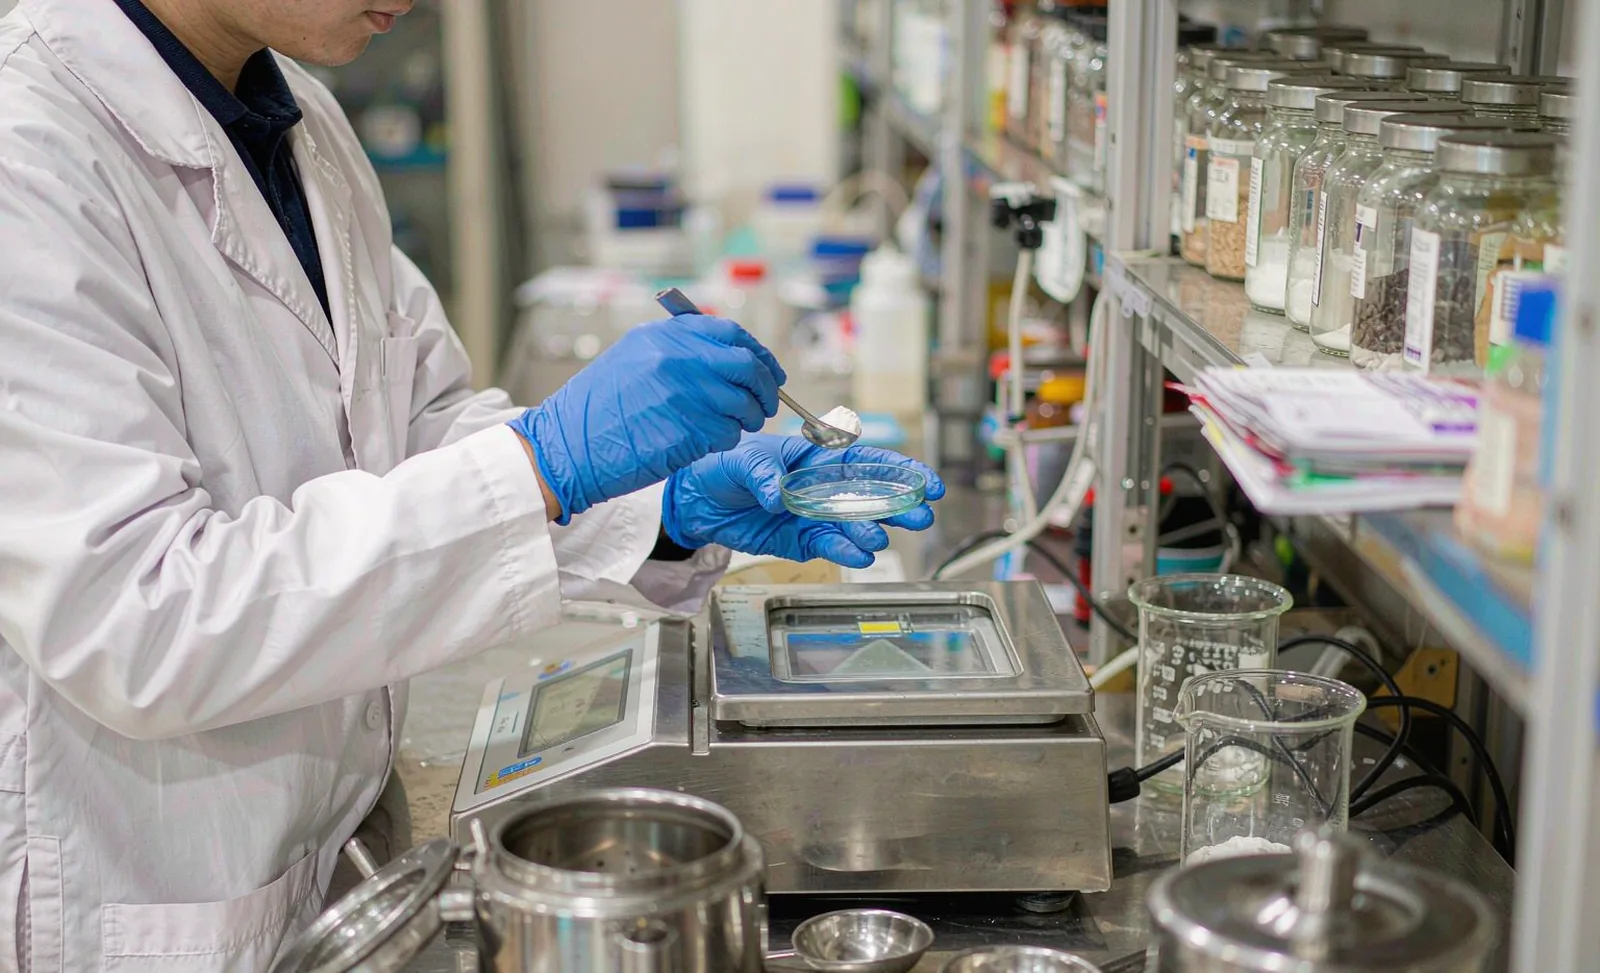
A lab or bench sample (often 0.5–5 kg of blended material, or a few hundred capsules) answers formulation questions: taste, dispersibility, capsule fill weight feasibility, basic blend feasibility with your excipient system, and preliminary assay of the blend. It does not answer scale questions: whether a 1,500-liter ribbon blender will achieve uniformity with this particle-size combination, whether the PS powder will flood or starve the feeder on a high-speed capsule filler, or whether drying temperatures in a fluid bed will push peroxide value upward. Write the sample stage's conclusions down as a bounded claim list ("1 kg scale blend uniformity RSD < 5% with this excipient ratio") rather than a general approval. This habit prevents the most common scale-up mistake: treating an R&D sample pass as an approval of the commercial process.
Stage 1: Pilot Batch at Intermediate Scale
The pilot batch — typically 10–20% of commercial scale, or at minimum the smallest batch your contract manufacturer's equipment can run meaningfully — is where ingredient personality meets real equipment. For PS products, the pilot should verify:
- Blend uniformity — sample the blend at defined time points and locations; PS at low inclusion rates (often 2–10% of a blend) makes uniformity genuinely challenging. Establish the mixing end-point with data, not a fixed clock time.
- Flow and weight control — for capsules and tablets, record bulk/tapped density, angle of repose if used, filler weight variation, and, for tablets, compression force windows and ejection behavior. PS's phospholipid nature can behave like a built-in lubricant at some levels and a friction problem at others.
- Thermal and shear exposure — confirm that processing temperatures and residence times (drying, conching for gummies, high-shear granulation) keep peroxide value and color within specification. Oxidation is PS's most realistic process risk.
- Moisture pickup — PS powders are hygroscopic to a degree; record humidity conditions during the pilot and whether open-handling steps need environmental control at scale.
Pilot output is a written process description with real parameter ranges — this becomes the master batch record's backbone.
Stage 2: Process Qualification and the First-Article Lot
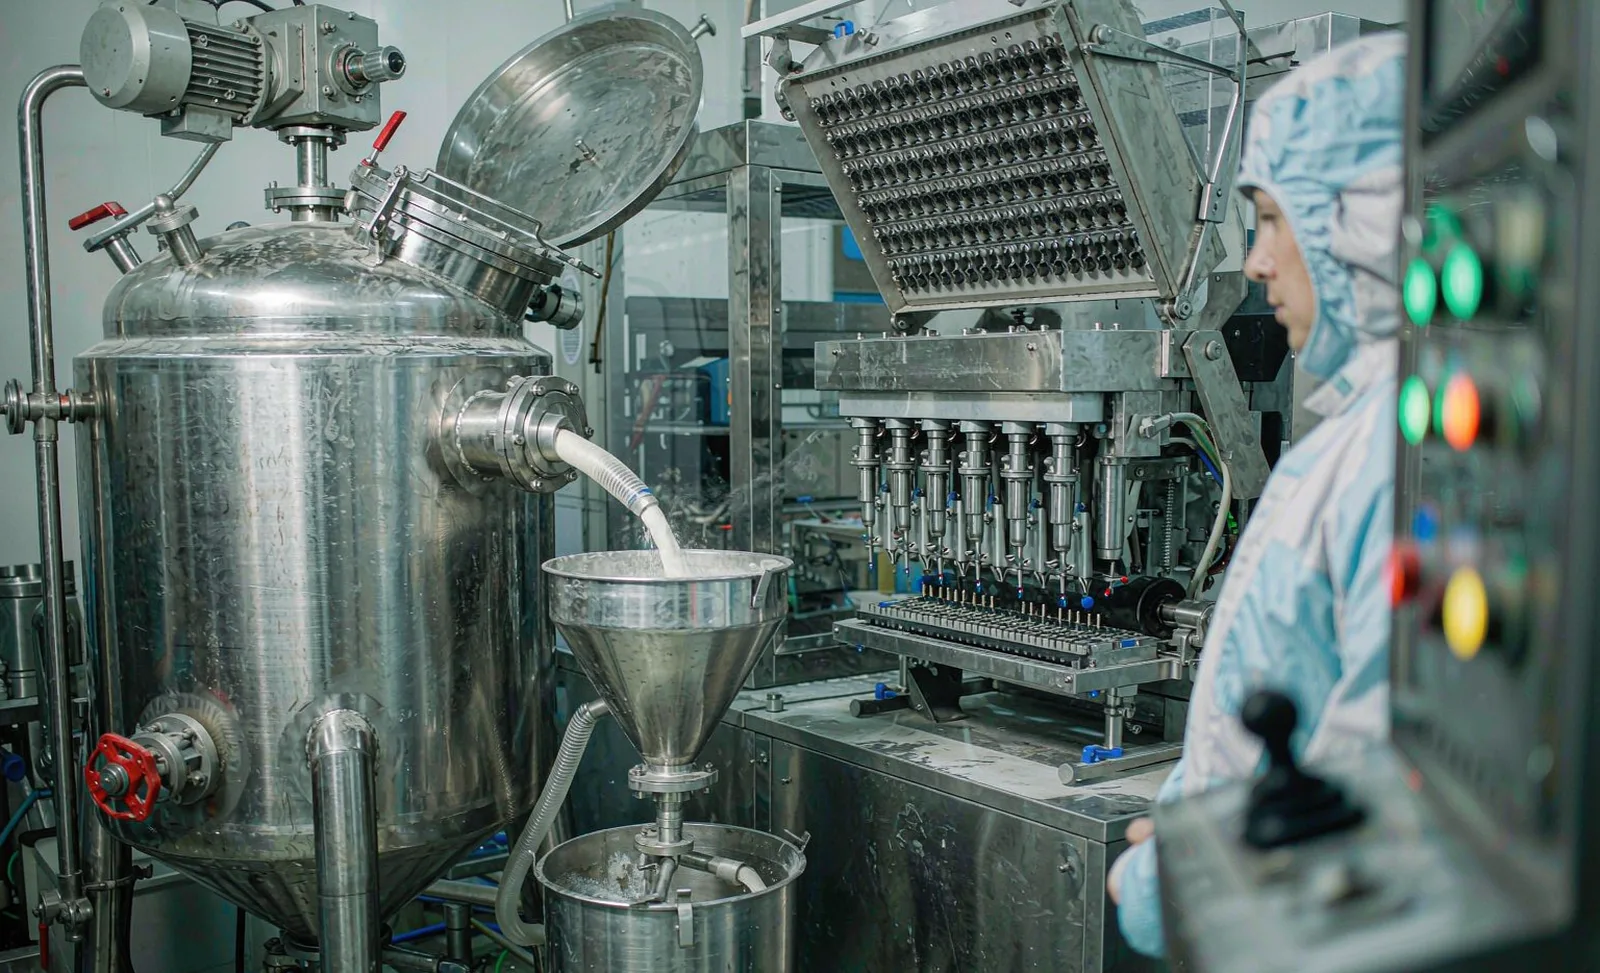
The first commercial-scale lot should be planned as a first-article / process-qualification batch, not a saleable production run. Its purpose is to demonstrate that the transferred process, at full scale, with full-rate equipment, delivers the finished specification. Practical requirements:
- Enhanced in-process sampling — blend uniformity at the commercial mixer scale, weight/fill variation over time, hardness and friability profiles for tablets, and moisture through any drying step.
- Full finished-goods release testing plus assay verification against the label-claim calculation built from the actual PS lot's assay and its correct basis (as-is versus anhydrous; total phospholipid versus PS fraction).
- Stability placement — the first-article lot, not a later lot, should seed the finished-product stability program, because its exposure represents the validated process.
- Documented deviation review — any parameter that required operator intervention or missed its target range is either fixed and re-demonstrated or formally justified.
FDA's process-validation guidance (Process Validation: General Principles and Practices) frames this as stages of process design, qualification, and continued verification — the same staged logic applies to dietary supplements even when a formal Stage 3 statistical program is not practical for low-volume SKUs. Align the contract manufacturer's approach with your quality agreement: define what "qualified process" means for this product before the first-article lot runs.
Stage 3: What Must Scale With the Batch — Documents and Calculations
Scale-up multiplies paperwork as surely as it multiplies powder. Items that must be regenerated at each stage, not copy-pasted:
- Master batch record with commercial-scale equipment, parameter ranges, and in-process checkpoints.
- Batch calculations using the actual PS assay of the commercial lot, including overage policy — decide once whether the label claim is met at end of shelf life and document the overage logic.
- Cleaning validation bridge — PS is a lipid-rich residue; confirm the CM's cleaning verification covers it (visual plus assay or TOC swabs) for shared equipment.
- Specification alignment — the finished-goods spec used for sample evaluation must be identical to the commercial release spec; silent edits here create retrospective compliance problems.
- Traceability file — one-step-back/one-step-forward traceability for the PS lot, the excipients, and packaging components, ready for both US recall-simulation drills and EU traceability expectations.
Managing PS-Specific Scale-Up Risks
Three ingredient-specific risks deserve their own controls. Oxidation during processing: cap open-handling time, avoid high-temperature drying, and verify peroxide value at the blend stage as well as on the incoming raw material. Segregation in transport and storage within the plant: low-inclusion PS powders can demix in conveying; define minimum transfer distances and verify uniformity at the point of fill, not only in the mixer. Assay method portability: confirm the HPLC method used at the CM matches the method behind the PS supplier's COA, or run a method-agreement comparison once during qualification. Each of these has sunk real launches; each costs one pilot-batch data set to control.
Sequencing and Timeline: A Realistic Sample-to-Bulk Plan
A disciplined pathway for a PS capsule or powder product looks like this: sample evaluation and formula lock (2–4 weeks), pilot batch and analysis (2–4 weeks including review), first-article commercial lot and enhanced testing (3–6 weeks including release testing), and stability placement with initial time-point data before full commercial launch (another 4–8 weeks for accelerated early reads). Total: roughly a quarter from locked formula to launch-ready, with each gate's documentation completed before the next begins. Programs that compress this by skipping the pilot almost always pay the time back — with interest — in first-article deviations, blend reworks, or a stalled release test. For EU-bound products, remember the finished-product documentation and labeling must also be finalized before commercial distribution; ISO 22000-based food-safety systems (ISO 22000:2018) provide a recognized framework for the CM-side controls your customers and auditors will ask about.
Perguntas frequentes
Q1: Why can't I skip the pilot batch if the sample already passed?
A sample proves the formula; a pilot proves the process on real equipment at a scale where mixing time, heat, and flow behavior resemble production. Skipping the pilot moves all those unknowns into your first commercial lot, where a failure costs the most time and money.
Q2: What blend uniformity target should I set for a low-inclusion PS product?
Common practice is relative standard deviation below 5% across a stratified sampling plan, but the right target depends on your inclusion rate and analytical method variance. Define the sampling locations, number of samples, and acceptance criterion in the pilot protocol — not after the data comes back.
Q3: Should the first commercial batch be sold or held as a qualification batch?
Plan it as a first-article/process-qualification batch with enhanced sampling. If everything passes and stability placement is done, the lot can usually still be released for sale — but it must be produced and judged as a qualification lot, with wider in-process data than routine production.
Q4: Does PS need special cleaning validation at the contract manufacturer?
Yes — phospholipid residues are lipid-rich and can be harder to remove and detect than typical powder residues. Confirm the CM's cleaning verification includes the PS-containing products, with appropriate swab methods or TOC limits, especially on shared blenders, dryers, and fillers.
Q5: How much overage should I build into a PS formulation?
Overage is a quality-policy decision, not a default. Set it from a documented calculation: label claim target, assay variance of the raw material, process and analytical loss, and shelf-life stability data. Many brands target the label claim at end of shelf life with a modest overage justified by stability results rather than a fixed industry percentage.
Conclusão
Sample-to-bulk scale-up for phosphatidylserine products is a staged proof problem: the sample proves the formula, the pilot proves the process, and the first-article lot proves the system at commercial scale — with documents scaling at every step. Brands that define each gate's deliverables before starting, watch PS-specific risks (oxidation, segregation, assay basis), and resist the temptation to skip the pilot reach commercial production with releasable lots and a clean audit trail. Those that compress the pathway usually donate the saved weeks back, plus interest, in deviations and rework.
Fontes
- FDA — Process Validation: General Principles and Practices (Guidance for Industry): https://www.fda.gov/media/71071/download
- FDA — Dietary Supplements: Current Good Manufacturing Practice overview: https://www.fda.gov/food/dietary-supplements
- ISO 22000:2018 Food Safety Management Systems: https://www.iso.org/standard/71726.html
- USP — Dietary Supplements Compendial Standards: https://www.usp.org/standards
- ICH Q1A(R2) Stability Testing of New Drug Substances and Products (stability design reference): https://database.ich.org/sites/default/files/Q1A%28R2%29%20Guideline.pdf
Próximas etapas recomendadas
- Revise o Phosphatidylserine página do produto.
- Comparar Soy PS e Sunflower PS.
- Verifique prova de fabricação e Qualidade e P&D.
Entre em contato com o departamento de vendas para obter documentos do produto
Compartilhe preferência de fonte, aplicação, país e quantidade anual.
Contato de vendas